Get 1 Month Free Trial on DMS & LMS |
Document control that holds up when the inspector walks in.
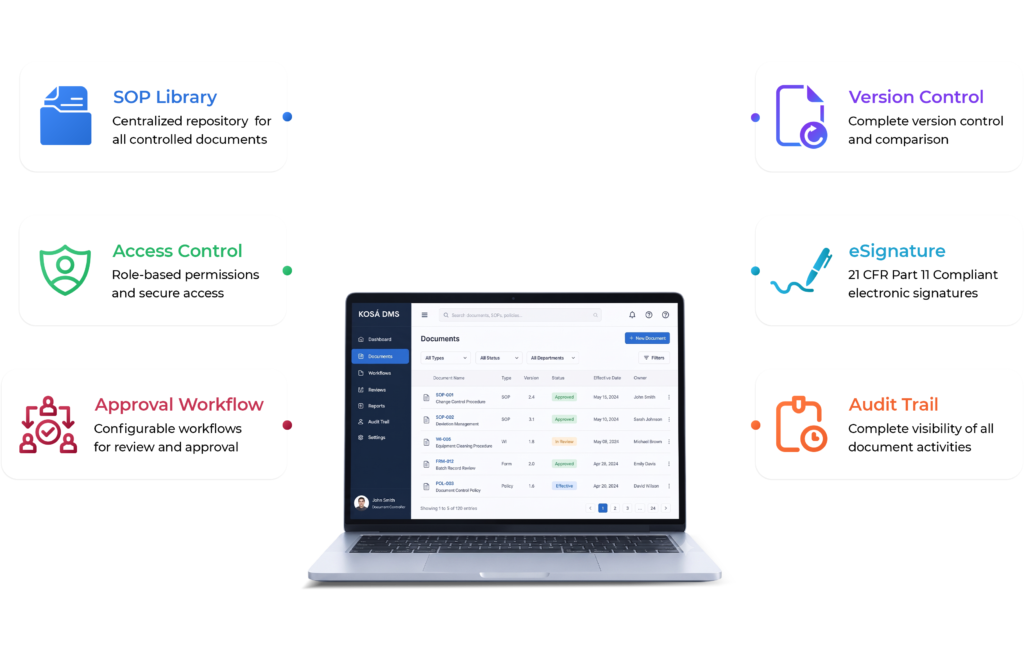
KOŚA DMS is the document management system for pharmaceutical industry teams that routes every SOP, batch record, and validation document through one configurable workflow -version-controlled, e-signed, and audit-trailed for 21 CFR Part 11, EU GMP Annex 11, and ISO 13485.
PharmaCorp
BioGenex
MediLabs
CuraVita
NovaPharm
What you can achieve
One controlled system replaces the shared drives, email threads and binders that quality teams stitch together by hand.
Centralise document control and replace manual routing with configurable workflows. Reduce manual coordination across teams.
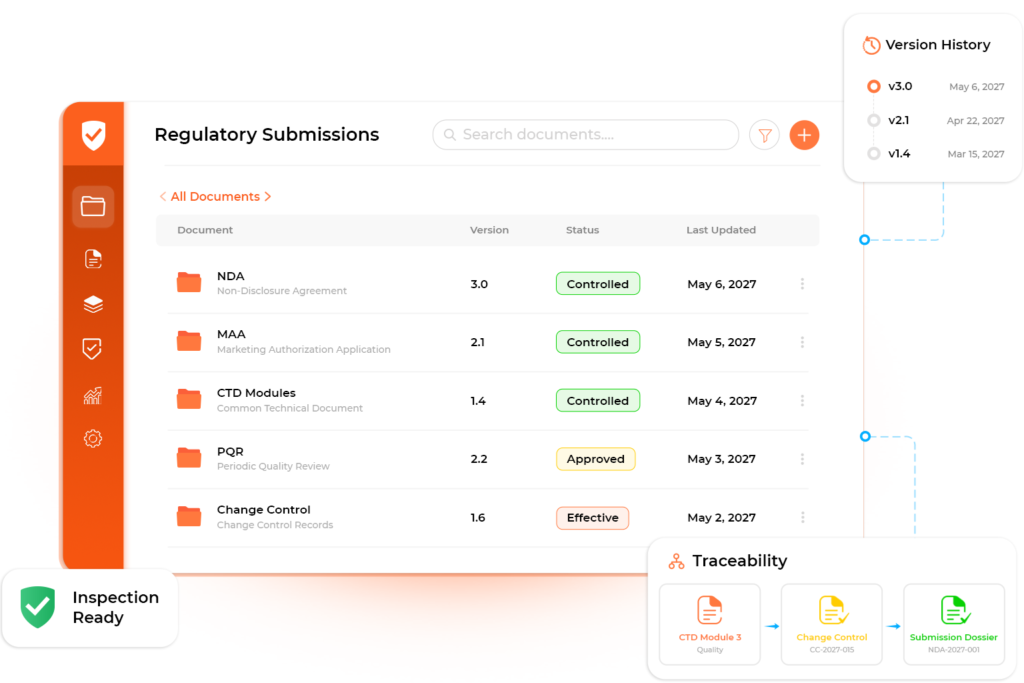
Designed to support FDA 21 CFR Part 11, EU GMP Annex 11 and ISO 13485 – keeping documentation inspection-ready at all times.
Monitor document activity, review backlogs and compliance KPIs in real time. Give QA leadership the visibility they need.
Go live in 6–8 weeks with structured implementation, training and 24/7 support – without disrupting existing operations.
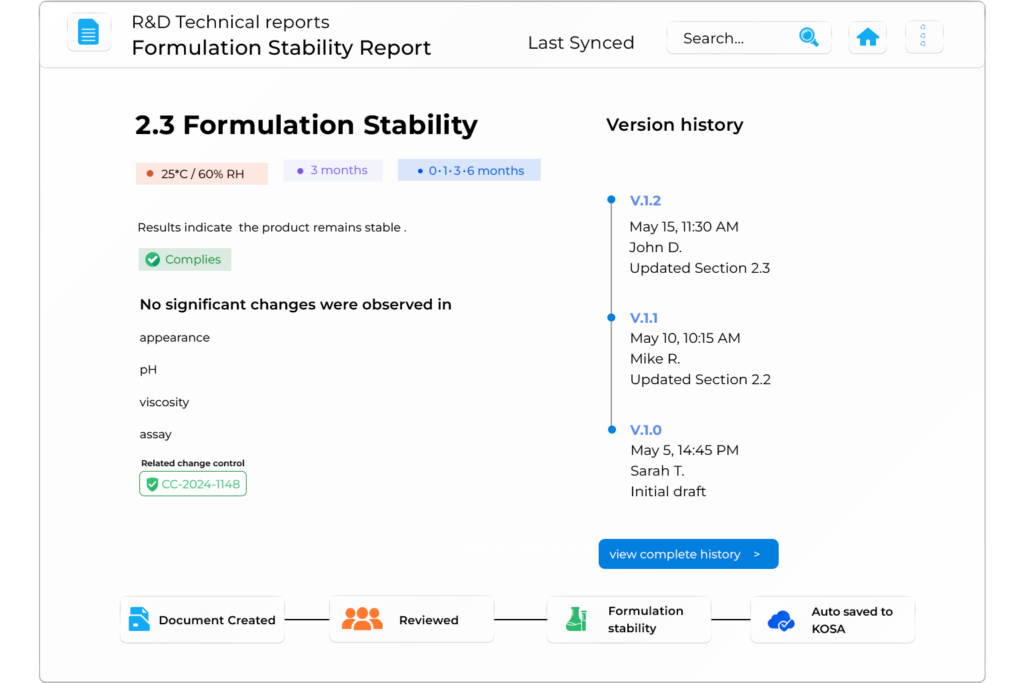
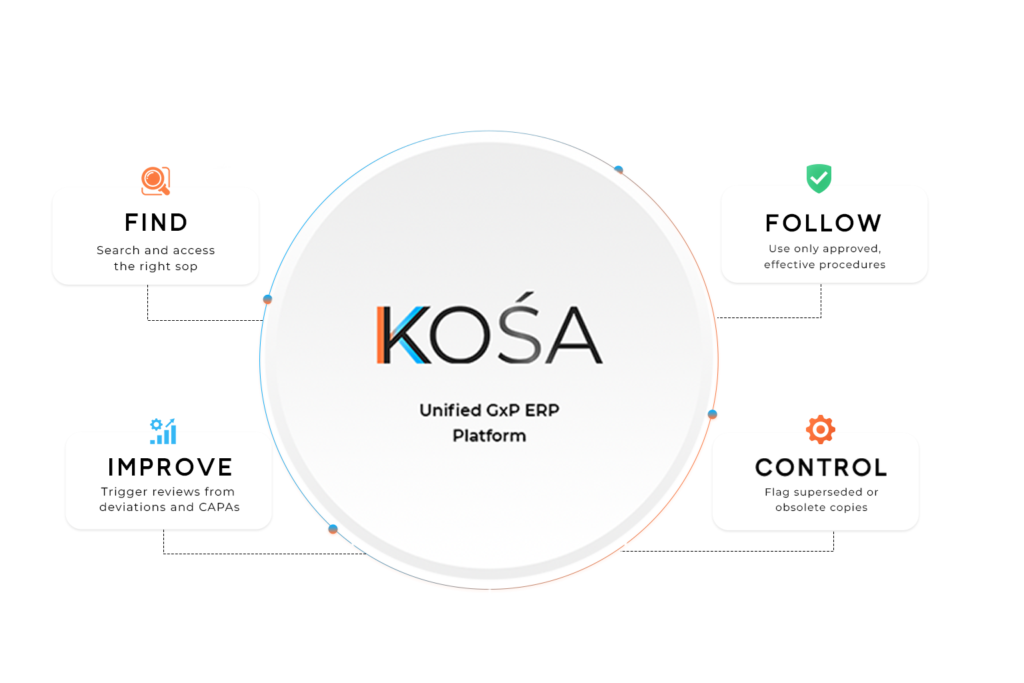
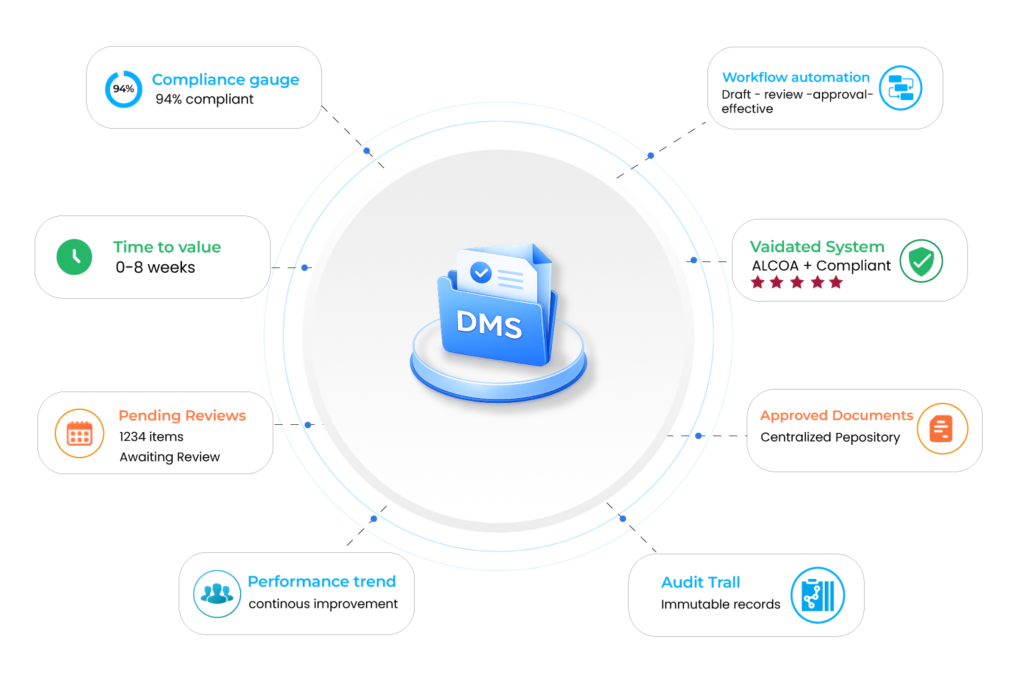
Why a centralized DMS matters
Electronic document management system for the pharmaceutical industry is critical to compliance, quality, and safety. Companies that meticulously organise and control their documents mitigate risk and maintain the highest quality standards across drug development and production.
Compliance
Good Documentation Practice (GMP) and GCP expectations are enforced inside the workflow itself – not reconciled afterwards at audit time.
Integrity
Every field, edit and signature is captured under ALCOA++ principles, so the record holds up on its own in front of an inspector.
Traceability
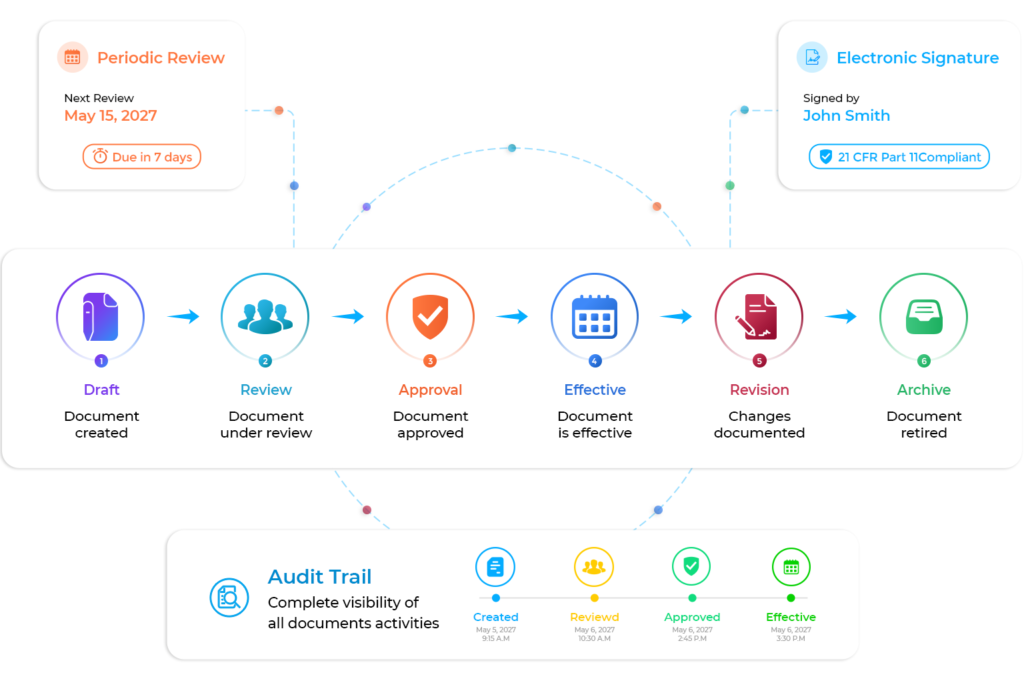
A single audit trail follows each pharma document management from the first draft through every review, approval, re-issue, and retirement.
Risk
Configuration drift, overdue reviews and unassessed changes are flagged while they’re still cheap to fix.
QA, regulatory, manufacturing and lab staff work from the same effective version in real time, wherever they sit.
Replaces the shared drives, email threads and binders that document control teams currently stitch together by hand.
What a pharma DMS controls
From raw-material receipt to product release, every step of the pharmaceutical document management system produces a controlled document. KOŚA pharmaceutical document management software is built around the records that matter most during an inspection.
Raw materials through release testing
Lab, cleaning and equipment SOPs
GCP design, methodology and results
NDA / MAA dossiers & correspondence
Raw material, intermediate and final QC
URS, IQ / OQ / PQ
Impact assessment and sign-off
PQR and stability data
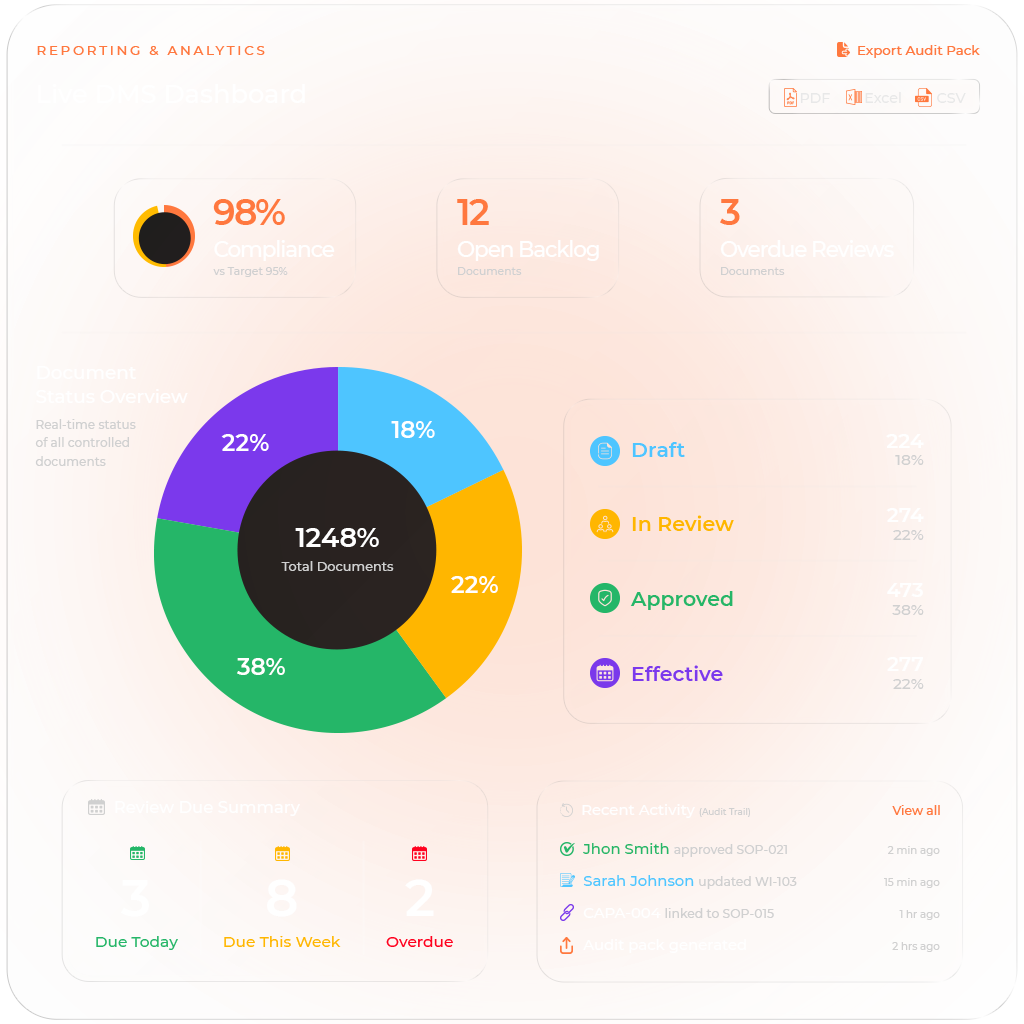
Reporting & Analytics
Live dashboards expose completion rates and compliance metrics – exports produce audit-ready documentation for every review.
Capabilities
Capability pillars that come standard – no add-ons, no surprises.
Draft, review, approval and effective-stage routing.
Traceable approvals aligned to 21 CFR Part 11.
Automatic change summaries for faster review.
The latest approved document with full traceability.
Control access by role, site or project.
Classify by type, process, owner and effective date.
Draft in Word while control stays inside KOŚA.
Full-text search and document properties.
Link documents to training assignments.
One consistent record across systems.
Track metrics, backlogs and compliance status.
Work simultaneously without losing version control.
Regulatory frameworks
KOŚA document management system for the pharmaceutical industry helps you implement document control processes that maintain traceability, controlled access, and documented oversight across the entire document lifecycle.
Audit trails, access control & signature binding.
Medical device design & document history records.
GMP-regulated manufacturing and quality activities.
Documentation, including technical file control.
Management for document repositories and access.
Technical file and post-market requirements.
Risk management and pharmaceutical quality systems.
Attributable, legible, contemporaneous, original, accurate.
Good documentation practice and data governance.
Risk-based validation, including IQ / OQ / PQ.
Documentation for medical devices.
Implementation
A phased, low-risk rollout designed around your validation package and audit calendar with 24/7 support from the first day.
Week 1–2
Document inventory, gap assessment and a structured cutover plan.
Week 3–4
Workflows, roles, templates and e-signature rules are configured to your SOPs.
Week 5-6
User acceptance testing alongside your validation package, ready for sign-off.
Week 7-8
Phased go-live with role-based training and 24/7 support from day one.
“Our document control team works much more effectively since we implemented KOŚA pharmaceutical document management software. The workflows are intuitive and our audits have never been smoother.”
Get started
Book a 30-minute walkthrough with a compliance architect – tailored to your SOPs, your workflows and your audit calendar.
FAQs
An electronic document management system built for 21 CFR Part 11 binds e-signatures to specific users and actions, timestamps every change in an immutable audit trail, and restricts access by role, so signature integrity and record traceability are enforced by the system itself rather than a manual SOP someone has to remember to follow.